This is the release for labs coming from qbase+. Bring your existing experiments across without losing data. Tag your samples the way you have always wanted to. And, for the first time, invite your team into shared workspaces and organizations so the work no longer lives on a single person's account.
With 2026.5, the qbase+ experiments you already have open in Clarida, and a lab can share its work in one workspace.
Welcome aboard.
Jan
New features
Bring your qbase+ experiments to Clarida
Migrating from one analysis tool to another usually means leaving years of work behind. With 2026.5 you can import your qbase+ experiments directly into Clarida and pick up where you left off. Sample annotations, assay definitions, plate layouts, run results, exclusion decisions, and analysis settings all come across.

Imported experiments open in a read-only mode that keeps everything you need usable: review the analysis, annotate samples, export the ELN workbook, and export results. The structure is locked; the analysis stays open to review and export. When you want to build new work on top, on a paid plan, duplicate it and make it your own.
Sample properties, all the way to charts
Tag your samples with the metadata that matters to your experiment: tissue type, treatment, timepoint, donor, dilution, anything you need. Properties are typed, so a numeric concentration sorts the way you would expect and a categorical group filters cleanly.

The properties do not stop at the table. In Data analysis you can now sort and group your relative quantification results by any property you defined. Rather than reading a long list of sample names, see your treatment groups side by side, your timepoints in order, or your replicates collapsed by donor. The visualization adapts to how your experiment is actually organized.

A cycling protocol and instrument per run
Cycling protocols and instrument choice now belong to the run, not the experiment. That matters when you run the same assay on two different instruments, or when one experiment spans a handful of runs with slightly different conditions. Each run carries its own protocol and instrument, and the run's record keeps both.

Previously, the design workflow derived the plate format for generating a run layout from the instrument set in the Cycling Protocol section. Plate format is now set in Run Layout, and the Cycling Protocol section has moved to the end, so you can set up each run's cycling to fit the plates laid out before it.

Run layouts based on source positions
The March release introduced generated run layouts whose structure you control. They work well if your assays and samples are stored in individual tubes and their relative positions do not matter. If your samples sat in strips and plates, or you wanted to control the order, your only option was manual run annotation. Source positions let you set where tubes and plates sit relative to each other, and the layout is still generated for you.

The result is a pipetting plan that matches what you can actually do at the bench, not a layout that looks tidy on screen but precludes time savings through the use of multi-channel pipets.
Storage holds experiments and libraries
Experiments, library samples, library assays, and tool outputs now live under a single Storage hub. Browse what you have, search across types, and open any item without losing your place in the rest of the app.

Work as a team in shared workspaces
Until now, a Clarida account was a one-person thing. With 2026.5 you can create a workspace, invite your colleagues, and share the work. Approve people who want to join with their institutional email, manage their roles, and remove access when someone leaves the lab. New members get a sign-in link in their invite email, with no password to set up before they can start.
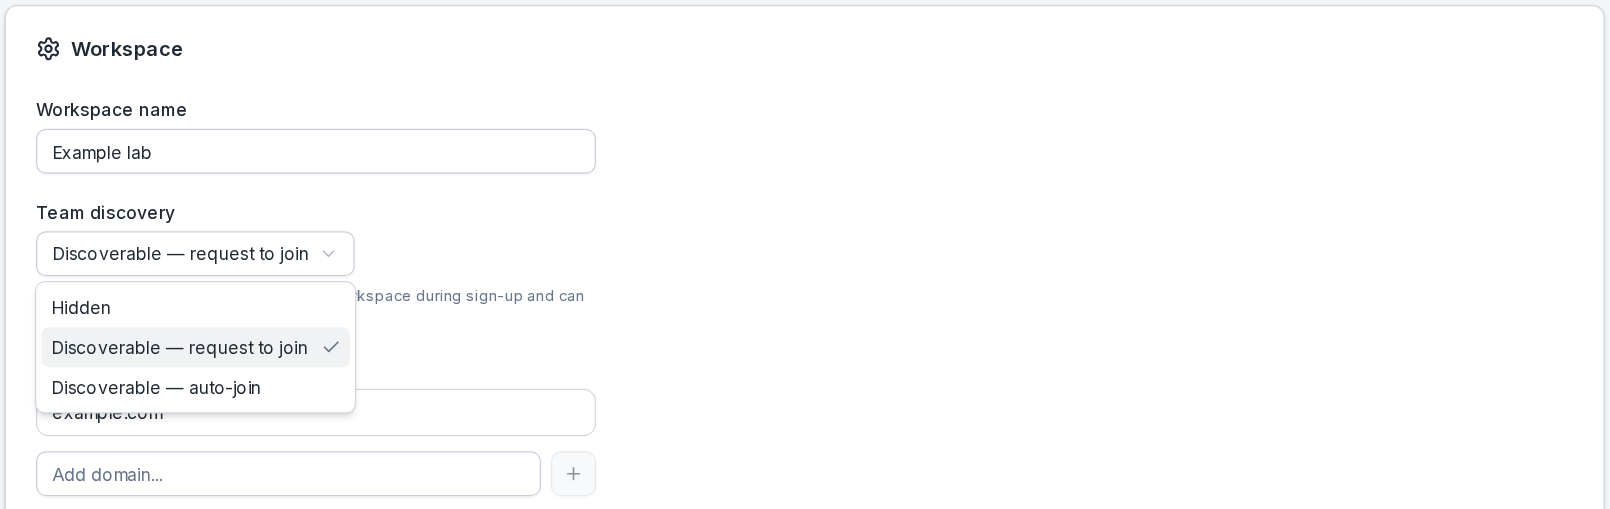

For larger setups (teaching labs, core facilities, partnerships), there is a layer above: organizations. An organization owns a shared usage pool that several workspaces can draw from, so a course coordinator can provision twenty student workspaces from one account once Clarida turns on several workspaces for the organization, and a PI can give their lab access without per-seat fragmentation. With experiment duplication, you can hand a new team member a fully populated example to work from.

Free accounts stay single-workspace; paid plans add collaboration. The free tier keeps the features it offered an individual researcher.
Quality of life
- Run Layout improvements · 384-well quadrant filling matches what robots and multi-channel pipets actually do, multi-plate spillover handles experiments larger than a single plate, and the plate visualization is improved to always reflect the current configuration.
- Reference gene finder PDF by email · Run an analysis on the public Reference gene finder and get a PDF report emailed to you. No account needed for the report itself; signing in to Clarida, optionally, keeps your work for later.
- NRQ ordering and grouping · Grouping in Data analysis can now respect your custom sample properties so you can present results the way your experiment is structured. Ordering of data by NRQ allows you to quickly sort samples by their expression level.
- Volumes to collect, per sample and assay · See exactly how much of each you need before you start pipetting.
- Aside settings on every section · Every workflow section with settings now has a settings button in the header to quickly jump to those settings in the aside panel. When jumping between sections, clicking anywhere in one now activates it so the corresponding settings are properly synced.
- Concentration popover stays open · The unit dropdown in the concentration popover no longer dismisses the popover when opened.
- Edited standards now update the results · Change a standard sample and efficiency, normalization, and quantification all recalculate automatically. See Changes that may affect you below.
- Feedback attachments · Attach more file types to feedback reports, and your contact information now auto-populates.
- Terms and privacy links at sign-in · Login page and invite/approval emails now link to the terms of service and privacy policy.
Patch 2026.5.1
A focused patch that makes sign-in more reliable at universities and companies with corporate email security, and fixes import errors for large plates and qbase+ workspaces.
- Sign-in code fallback for magic links · If your magic link doesn't work, common at institutions where Microsoft Defender, Mimecast, Proofpoint and similar scanners pre-click email links, every magic-link email now also includes a 6-digit sign-in code. When the link fails, look for "Enter your 6-digit code instead" on the login page, type the code from the email, and you're in.
- Sign in with Microsoft · At universities and pharma companies on Microsoft 365 / Azure AD, you can now sign in with your existing institutional account, available on the login page alongside Google. Federated SSO bypasses the email channel entirely, avoiding the prefetching issue above for organizations that support it.
- Support for large imports · Workspaces containing 1536-well plates (4-character coordinates like AF48) or qbase+ imports with serial well indices above 999 previously failed to save with a database error. They now import correctly.
Patch 2026.5.2
A targeted fix for the public Reference gene finder when running large miRNA-scale panels.
- Large geNorm panels no longer stall · Running geNorm on panels with hundreds of candidate genes (the typical miRNA discovery scenario, e.g. 376 genes × 16 samples) previously hung the analysis. The pairwise stability calculation has been rewritten to use vectorized math, and the same workload now completes in a few seconds. Smaller panels are unaffected; rankings and M values are bit-identical to before for the cases that already worked.
Changes that may affect you
Some results change. What to re-check is below.
Added on October 30, 2026: changes we left out at release.
A changed standard sample did not update the results. When you added, edited or deleted a standard, efficiency, normalization and quantification kept the old values until something else changed. They now recalculate at once; re-open experiments where you edited a standard.